
Score
erlotinib through the run: 29 heavy atoms, flexing in place.
Pose validity
- No ligand-protein heavy-atom contact under 2.2 A pass
- Ligand bond lengths within 25 % of the ideal pass
- Stereocentres unchanged from the input SMILES pass
- Ligand centroid inside the docking box pass
Predicted affinity
| Docked pose | After MD, 5 frames | Change | |
|---|---|---|---|
| pIC50 | 6.53 | 6.46 ± 0.02 | -0.07 |
| Boltz-2 value | -0.53 | -0.46 ± 0.02 | +0.07 |
| Binder probability | 0.52 | 0.51 ± 0.01 | -0.01 |
Predicted pIC50 6.46 after MD, 0.07 log units weaker than the docked pose.
affinity_pred_value is log10(IC50) with IC50 in micromolar, lower is stronger; pIC50 = 6 - affinity_pred_value, higher is stronger.
Boltz-2's affinity head. The complex was held within 2 A of the structure this pipeline produced while Boltz-2 rebuilt it, so the head reads the docked and relaxed conformations rather than one of its own. It is not a free energy, it is not comparable across targets, and it is not evidence that the pose is correct: nothing here is weighed against a crystal, so none of it enters the score above.
Pocket backbone through MD: 0.65 → 0.98 Å from the crystal, so MD moved it 0.33 Å further away. Reported, not scored: in every run measured so far MD has moved the pocket away, so this says what happened rather than how well it went.
Complex
Overlay
Motion
The production run in full: 50 frames over 490 ps. Drag to turn, scroll to zoom, Play to run the trajectory.
Docked poses
The top pose is 1.59 A from the crystal ligand: the search found it and the scoring function agreed.
Pose ranking
| # | Score | Vina | pEC50 | Vinardo | RMSD | Centre | Shape | Contact |
|---|---|---|---|---|---|---|---|---|
| 1 | -16.47 | — | — | -7.59 | 1.59 | 0.74 | 1.38 | 3.19 |
| 9 | -15.84 | — | — | -7.36 | 3.35 | 0.85 | 2.31 | 3.19 |
| 6 | -15.99 | — | — | -7.23 | 8.64 | 3.49 | 2.27 | 3.01 |
| 8 | -15.87 | — | — | -6.83 | 4.23 | 0.87 | 2.38 | 2.80 |
| 4 | -16.24 | — | — | -6.65 | 8.22 | 0.64 | 2.08 | 3.08 |
| 7 | -15.98 | — | — | -6.47 | 8.46 | 0.90 | 1.70 | 3.24 |
| 5 | -16.02 | — | — | -6.43 | 8.24 | 0.95 | 1.25 | 3.32 |
| 3 | -16.24 | — | — | -5.86 | 8.44 | 1.42 | 2.47 | 3.04 |
| 10 | -15.83 | — | — | -5.86 | 7.22 | 2.74 | 2.53 | 3.22 |
| 2 | -16.32 | — | — | -5.46 | 8.74 | 1.03 | 2.39 | 2.95 |
Score and Vina in kcal/mol; RMSD, Centre, Shape and Contact in A. RMSD is to the crystal ligand where it lies. Centre is the distance between the two centroids: it says whether the pose is in the right pocket at all. Shape is the RMSD after superposing the two ligands freely, which throws the placement away and asks only whether the molecule is folded the same. A low Centre and a low Shape beside a high RMSD is a ligand in the right site, turned the wrong way round. Contact is the closest heavy-atom approach to the receptor; under 2.2 A is a clash. Pose 1 went to MD; a highlighted row below it is the pose nearest the crystal.
Vinardo re-scored these same ten coordinates and decides which pose goes forward. It sampled nothing and moved nothing; it chose. Measured over nine pose sets, the docking engine put a pose within 2 Å of the crystal first once and this function did it four times, and on the run the engine got right it picked the same pose. Here the two agree, which is two independent functions on the same answer.
Boltz-2 scored pose 1 alone, before and after MD: see the affinity block in the Score panel. Running the affinity head on all 10 poses would cost about 60 minutes and is not done by default.
Run
- ProteinP00533
- Liganderlotinib
- Familykinase
- Reference1M17 · AQ4
- TM-score, whole chain0.958
- Pocket Ca atoms fitted51
- Analysis30.3 s
Engine versions
- gobsmacked_run1.0.0
- python3.11.16
- platformDarwin arm64
- rdkit2026.03.1
- openmm8.6
- mdtraj1.11.1
- torch2.10.0
- pdbfixerinstalled
- openmmforcefields0.16.0
- pandadockv4.1.1
- The receptor was co-folded with the ligand and the predicted ligand pose was then discarded, so docking starts from a pocket shaped around this ligand rather than an apo one. The discarded pose is in cofold/cofold_ligand.sdf.
- The docking engine ranked its own pose 1 at position 1 of 10. A second scoring function (vinardo) placed it first, and it is the pose carried forward. Both sets of scores are in poses/, and the engine's original rank is a column in scores.csv.
- The docking box was re-centred by 48.1 A during the run, from [-9.87, 33.08, 14.14] to [4.56, -11.34, 2.73], to put it in the frame of the receptor this run built. The campaign recorded here is the one that ran.
Most displaced pocket side chains
| Residue | Side-chain RMSD | chi1 model | chi1 crystal | difference |
|---|---|---|---|---|
| Glu780 | 3.26 A | -68.1 | 89.9 | 158.0 |
| Gln767 | 2.84 A | -165.8 | 162.9 | 31.4 |
| Glu722 | 2.81 A | -175.8 | 62.5 | 121.7 |
| Gln763 | 2.57 A | -38.1 | -56.1 | 18.0 |
| Asp776 | 2.55 A | -56.8 | -70.9 | 14.2 |
| Ser696 | 2.5 A | 67.9 | -58.0 | 125.9 |
| Thr766 | 2.25 A | -161.2 | 50.7 | 148.2 |
| Phe699 | 2.19 A | 87.6 | 54.2 | 33.4 |
| Phe832 | 2.12 A | -41.3 | -65.7 | 24.4 |
| Thr701 | 2.03 A | -25.8 | 59.7 | 85.6 |
Interactions
| Residue, 1M17 numbering | MD final | Crystal |
|---|---|---|
| 694 | hydrophobic | hydrophobic |
| 702 | hydrophobic | — |
| 721 | hydrophobic | hydrophobic |
| 764 | hydrophobic | hydrophobic |
| 766 | hydrophobic | hydrophobic |
| 768 | — | hydrophobic |
| 769 | hbond | hbond |
| 820 | hydrophobic | — |
Residues are numbered as the crystal numbers them, because that is the only
numbering both structures share. The PandaMap panel above uses the complex's own.
Jaccard overlap with the crystal: pose 1
0.714, MD final
0.625.
Interaction map
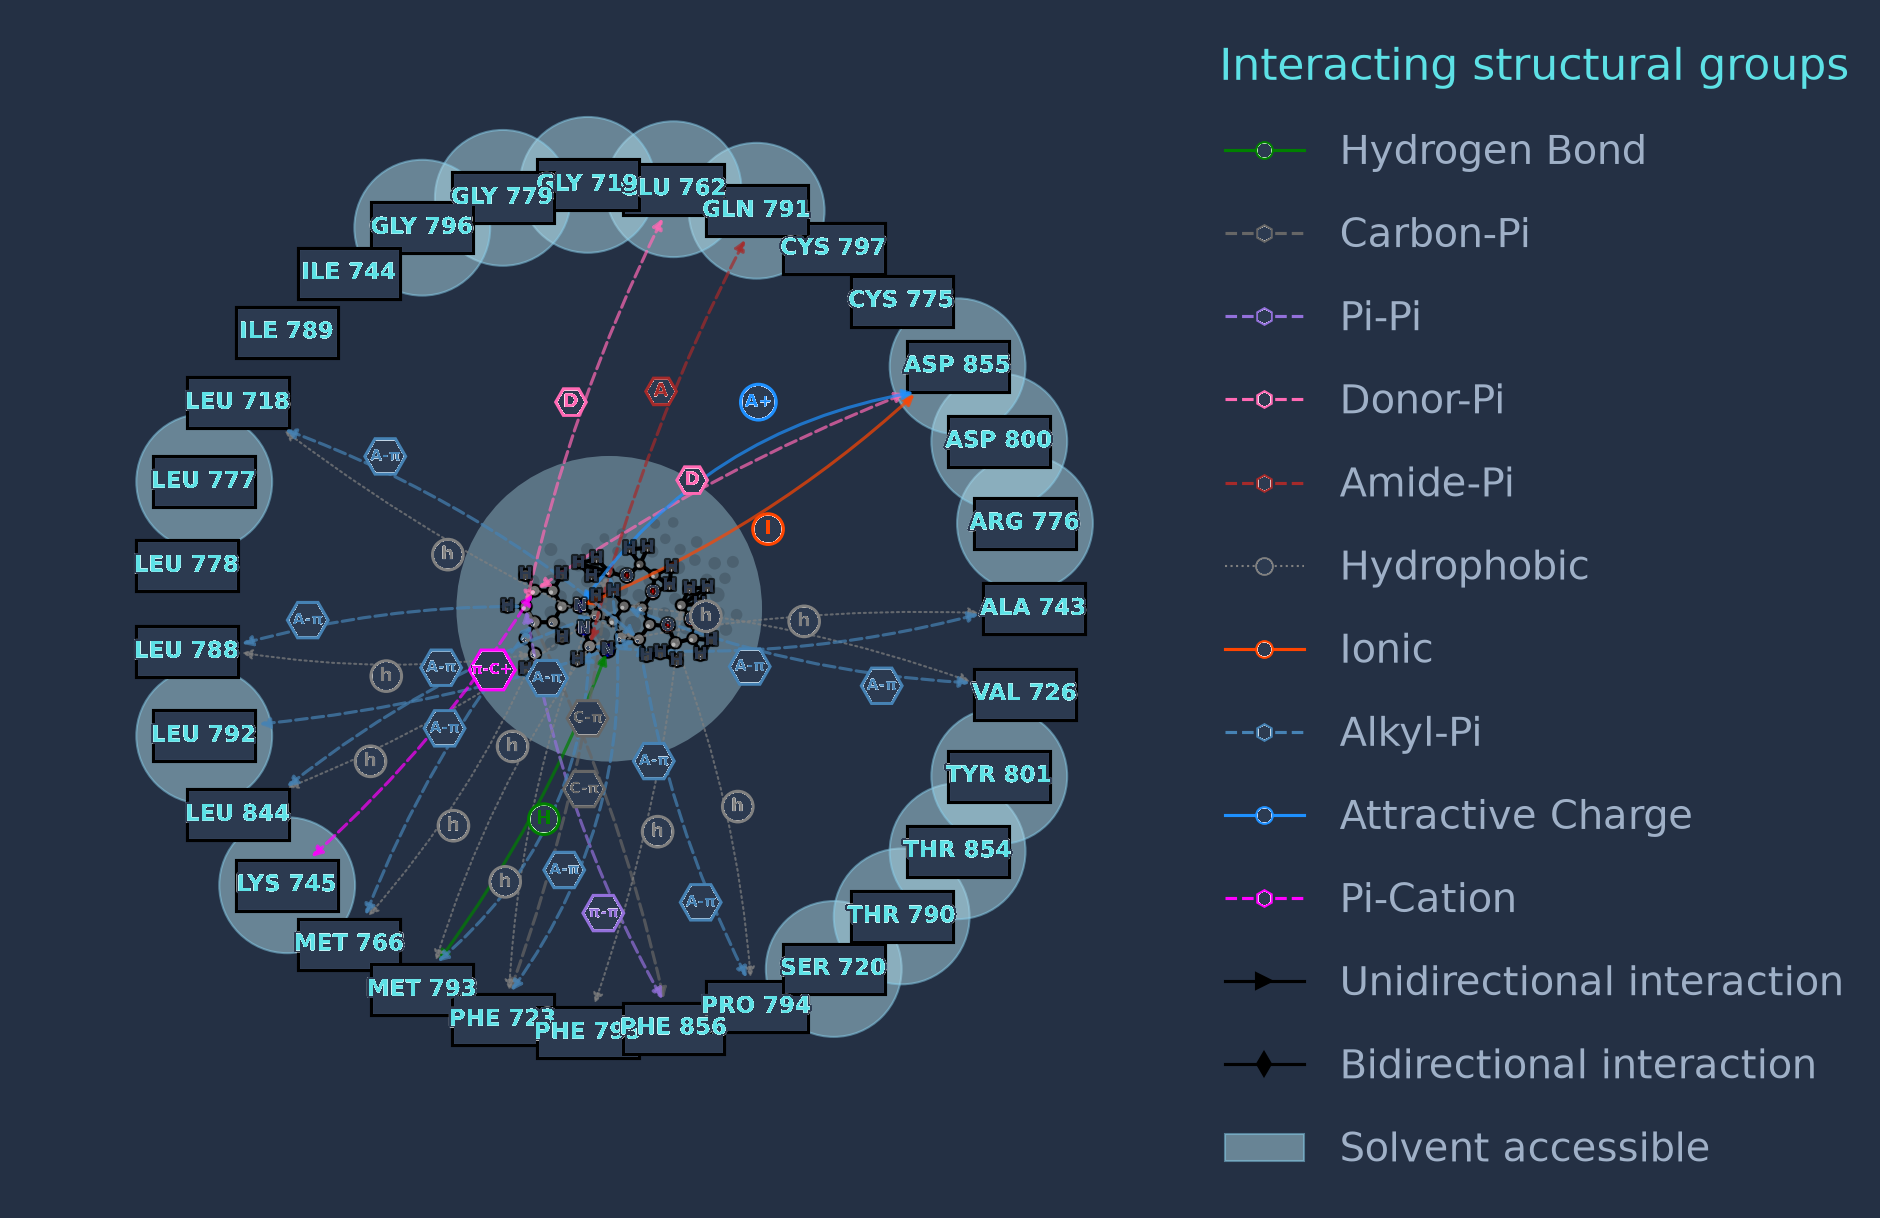
PandaMap's empirical estimate: -0.53 kcal/mol (Very weak / no binding expected). Empirical estimate ±2–3 kcal/mol; not a substitute for FEP/MM-GBSA. Hover any residue or legend entry for what it means.
Mode: kinase switches
- DFGin
- alphaC helixin
- Hinge H-bonds1
- Gatekeeper contactThr790
- Back pocket I occupied
- Back pocket II empty
- Front pocket occupied
- Adenine pocket occupied
- Solvent exposed 0% of ligand
Dynamics
Report
A printable page with the scorecard, the mode verdict, the interaction table and the key traces. Printing preserves the panel colours.